
Natural Product–Ribosome Structure Enables Design of New Anti-TB Drug Leads

Tuberculosis (TB) is one of the world’s leading causes of death from a single infectious agent, second to human immunodeficiency virus (HIV). However, TB-HIV co-infection is a serious problem because current TB drugs (such as rifampin) induce hepatic metabolic enzymes (such as CYP2C9 and CYP3A4) that degrade anti-retroviral drugs, which in turn renders patients in an immunocompromised state that accelerates the progression of their TB.
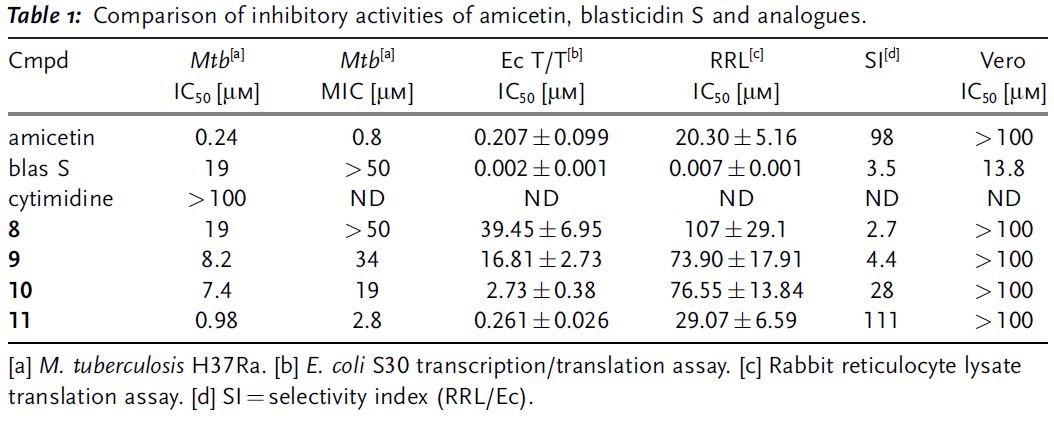
In their search for new anti-TB drug leads that do not induce CYP activation, the Looper group and collaborators identified the known natural product amicetin (Figure 1) and found that it possesses potent anti-TB activity (IC50 = 0.24 µm), but less than 3-fold induction of CYP3A4 and CYP2D6 (rifampin induction levels are 47- and 6-fold, respectively). Therefore, the group decided to utilize to modify the pharmacophore of this natural product to address the need for new TB therapies. Although amicetin and its natural relatives have been shown to inhibit protein synthesis by binding to the prokaryotic ribosomal P-site, the molecular nature of these interactions have not been fully elucidated. The group co-crystallized purified amicetin with the 70S subunit of the ribosome of Thermus thermophilus (Tth) and generated a crystal structure at 3.5 A resolution (Figure 2).
On the basis of these findings, the authors were able to hypothesize that the acid labile cytosine glycosidic linkage could be removed, since its hydrolysis products cytimidine and cytosamine are both inactive, and the disaccharide moiety could be simplified such that the synthetic chemistry is amenable to basic medicinal chemistry tactics. Their synthetic approach is summarized in Scheme 1. Notably, the removal of the disaccharide significantly simplifies the synthetic route, and the only chiral information in the synthetic analogues is embedded in commercially available α-methylserine.

Remarkably, compound 11 is only ~4 fold less active than amicetin, despite the removal of 8 stereogenic centers. This analogue also maintained selectivity for inhibiting bacterial (vs. eukaryotic) translation, in contrast to blasticidin S (Table 1). Remarkably, no cross-resistance was observed, and 11 maintains activity against TB strains resistant to rifampicin (RIFR), isoniazid (INHR), and levofloxacin (FQR), as shown in Table 2. The activity of these analogues is limited to TB and some gram positive bacterial strains, although the group is currently working on extending the spectrum of activity of these compounds.
This study provides the first structural characterization of ribosomal P-site inhibitor that selectively inhibits prokaryotic translation. In addition, the study demonstrated that structurally simplified analogues can provide tractable leads for new anti-TB therapies.
Reference:
Serrano, C. M.; Kanna-Reddy, H. R.; Eiler, D.; Koch, M.; Tresco, B. I. C.; Barrows, L. R.; VanderLinden, R. T.; Testa, C. A.; Sebahar, P. R.; Looper, R. E. Angew. Chem. Int. Ed., 2020 (Just Accepted) doi: 10.1002/anie.202003094.

Comments

Popular posts from this blog

Lysine-Targeted Covalent Inhibitors Gaining Traction

In the past decade, targeted covalent inhibitors (TCIs) has experienced a resurgence due to its sustained target engagement and improved proteome-wide selectivity. Currently, targeting cysteine residues with acrylamides and other α, β-unsaturated carbonyl compounds is the predominant strategy in TCI development. Although lysine is more prevalent than cysteine and possesses lower mutation rates, lysine has been much less frequently considered and more challenging as a residue targeted by TCIs due to its low nucleophilicity. Recently, Jack Taunton and his colleagues reveled their two related work in JACS .   Based on the co-crystal structure of Hsp90 with a reversible inhibitor PU-H71, they designed two arylsulfonyl fluorides 1 and 2 and co-incubated them with Hsp90 protein (Figure 1a, b). Mass spectrometry detected the formation of a 1:1 covalent adduct. Mutation of Lys58 to arginine abrogated the covalent modification, indicating that the TCI selectively modified Lys58 (Fig...

First PROTAC Degradation of Mulit-Pass Transmembrane Protein

Cancer cells exploit proteins involved in energy metabolism to maintain a steady supply of building blocks necessary for increased proliferation. Many transporters play a role in the process of shuttling water, nutrients, ions, and various metabolic products across the cell membrane, including the solute carrier (SLC) group of proteins. The SLC protein group consists of more than 400 proteins organized into more than 50 families. For example, the SLC9 family is involved in intracellular pH homeostasis, with increased SL9 activity resulting in an elevated intracellular pH and cytosolic alkalinization in cancer. Currently there 12 FDA approved drugs targeting SLC proteins for conditions ranging from hypertension to depression, but recent studies suggest SLC proteins may be attractive targets for cancer drug development. With this end goal, Bensimon et al., aimed to investigate if multi-pass transmembrane proteins like SLCs can be chemically degraded. To first explore this aim, th...

Grafting Alkyne Handles on Cysteines

One of the most common ways to study biomolecules is to label specific amino acids, mostly cysteine, with chemical handles for further modification. Azide-alkyne click reaction is a very popular strategy due to its high reactivity and reaction rate. Therefore, numerous reagents have been developed to specifically label proteins with terminal alkynes, including iodoacetamide alkynes and maleimide derivatives. However, limitations like insufficient selectivity and low stability remain as long-lasting concerns. To address these problems, one common strategy is introducing a linker between the targeting residue and the alkyne moiety , which potentially brings in unwanted structural changes. A reagent that can introduce small-sized terminal alkyne with one step under mild conditions is in demand. Scheme 1 . Cysteine-labeling study in this work In this study, the Waser group presented a series of reagents that selectively ethynylate cysteine in a one-pot manner ( Scheme 1 ). The ...